Medical device quality cannot be evaluated only by examining the finished product. It also depends on the systems used to select materials, qualify vendors, control manufacturing, document changes, investigate complaints, and correct problems. A device may appear acceptable during a final inspection while still being produced through inconsistent or poorly controlled processes.
ISO 13485 certification helps medical device suppliers demonstrate that they operate a structured quality management system designed for the medical device industry. The standard establishes requirements covering responsibilities, documentation, risk-based controls, product realization, traceability, supplier management, complaint handling, and continual system maintenance.
This certification can be important for manufacturers of finished devices as well as organizations that provide components, packaging, sterilization, testing, calibration, distribution, and other services that influence device quality. Although the precise regulatory obligations depend on the organization’s role and the markets it serves, ISO 13485 has become one of the most widely recognized quality benchmarks in the medical device supply chain.
Its significance has increased in the United States. The FDA’s Quality Management System Regulation, or QMSR, became effective on February 2, 2026, and incorporates ISO 13485:2016 by reference as the foundation of the agency’s medical device current good manufacturing practice requirements. However, certification itself does not automatically establish compliance with every applicable law, replace FDA oversight, or authorize a medical device for sale.
For medical device suppliers, the value of ISO 13485 certification lies in what the underlying system can accomplish: more consistent operations, clearer accountability, stronger traceability, better risk control, and greater confidence among manufacturers, healthcare organizations, regulators, and other business partners.
Key Takeaways
- ISO 13485 is an internationally recognized quality management system standard developed specifically for medical device organizations.
- Certification provides independent evidence that a supplier’s quality management system has been assessed against the standard’s applicable requirements.
- ISO 13485 certification is not the same as FDA registration, device clearance, approval, or authorization to market a product.
- The standard helps suppliers control documentation, purchasing, production, traceability, complaints, nonconformities, and corrective actions.
- Certification can make supplier qualification more efficient because customers receive a recognized form of quality-system evidence.
- A well-maintained ISO 13485 system can reduce preventable errors, uncontrolled changes, inconsistent records, and supply-chain risk.
- ISO 13485 supports international market readiness, but suppliers must still meet the laws and customer requirements applicable to each country.
- Certification is most valuable when the quality management system is actively used in daily operations rather than treated as an audit exercise.
What ISO 13485 Certification Means For A Medical Device Supplier
ISO 13485:2016 is the current edition of the international standard titled Medical devices—Quality management systems—Requirements for regulatory purposes. The International Organization for Standardization confirmed the standard as current following its 2025 review. It is designed for organizations involved in one or more stages of a medical device’s life cycle, including design, production, storage, distribution, installation, servicing, and related support activities.
A Medical Device-Specific Quality Framework
ISO 13485 is different from a general quality standard because it focuses on the controls needed in a regulated medical device environment. Its requirements reflect the importance of patient safety, documented evidence, risk management, traceability, process validation, and regulatory compliance.
Depending on its activities, a supplier’s quality system may need to address:
- Quality policies and measurable objectives
- Management responsibility and periodic management reviews
- Employee competence and training
- Document and record control
- Customer and regulatory requirements
- Design and development controls, when applicable
- Purchasing and supplier controls
- Production and service processes
- Product identification and traceability
- Monitoring and measuring equipment
- Validation of processes that cannot be fully verified afterward
- Complaint handling and regulatory reporting support
- Control of nonconforming products
- Corrective and preventive action
- Internal audits and quality-system monitoring
Not every requirement necessarily applies to every organization. The scope depends on the supplier’s responsibilities, activities, products, services, and applicable regulatory requirements. Any justified non-applicability must be handled carefully and documented within the boundaries permitted by the standard.
Certification Involves Independent Assessment
An organization becomes certified after an external certification body audits its quality management system and determines that it conforms to the applicable ISO 13485 requirements. Certification normally involves an initial assessment followed by surveillance audits and periodic recertification.
Customers should examine more than the existence of a certificate. Relevant details include:
- The legal entity named on the certificate
- The physical sites included
- The products, services, and activities within the certified scope
- The certificate’s issue and expiration dates
- The certification body
- Any exclusions or limitations
- The certificate’s current status
A supplier may hold a valid certificate that covers one manufacturing site or service but not another. Buyers should confirm that the scope aligns with the work the supplier will actually perform.
It Is A System Certification, Not A Product Approval
ISO 13485 certification applies to an organization’s quality management system. It does not certify that every product is safe, effective, FDA-cleared, FDA-approved, or legally marketable in every country.
That distinction is critical. Device authorization and quality-system certification serve different purposes. A company must still determine and satisfy the classification, registration, listing, premarket, labeling, reporting, and other obligations applicable to its products and target markets.
How ISO 13485 Supports Regulatory Readiness
One of the strongest reasons for medical device suppliers to pursue ISO 13485 certification is its connection to regulatory quality-system expectations. Many jurisdictions either recognize the standard, rely on it within their conformity-assessment structures, or use requirements closely aligned with it.
Alignment With The FDA’s QMSR
The FDA’s Quality Management System Regulation became effective on February 2, 2026. The regulation amended 21 CFR Part 820 and incorporates ISO 13485:2016 by reference, along with certain terminology from ISO 9000:2015.
The change brings the U.S. device quality framework into closer alignment with internationally recognized quality-management principles. The FDA explains that the QMSR applies to finished device manufacturers intending to commercially distribute medical devices, subject to defined exemptions and other provisions.
This development makes familiarity with ISO 13485 especially important for suppliers serving the U.S. market. Nevertheless, certification does not equal FDA compliance. U.S. manufacturers must comply with the Federal Food, Drug, and Cosmetic Act and all applicable FDA regulations. Where a conflict exists, U.S. law controls. FDA-specific requirements involving areas such as labeling, complaint records, adverse-event reporting, corrections and removals, and Unique Device Identification may also apply.
A supplier that is not directly subject to every QMSR requirement may still be expected by a finished-device manufacturer to maintain comparable controls. The manufacturer remains responsible for ensuring that purchased products and services meet specified requirements.
Supporting European Market Requirements
Under the European Union Medical Device Regulation and In Vitro Diagnostic Medical Device Regulation, manufacturers must establish, document, implement, maintain, update, and continually improve an appropriate quality management system. Harmonized standards may help organizations demonstrate conformity with relevant legal requirements, although the regulations themselves remain controlling.
ISO 13485 certification can provide useful evidence of a structured quality system when a supplier works with manufacturers placing devices on the European market. It may also support interactions with notified bodies and customer audits. However, certification alone does not grant CE marking or demonstrate compliance with every requirement of the applicable EU regulation.
Preparing For Multijurisdictional Audits
Organizations supplying multiple countries can face overlapping expectations. One customer may require FDA-oriented controls, another may focus on EU conformity assessment, and another may participate in the Medical Device Single Audit Program.
ISO 13485 gives suppliers a common foundation on which jurisdiction-specific procedures can be built. This can reduce fragmented documentation and help teams identify which requirements apply globally and which require local supplements.
The Medical Device Single Audit Program, for example, permits recognized auditing organizations to conduct one audit that addresses participating regulatory authorities’ applicable requirements. ISO 13485 forms an important basis for that auditing model, but participation and acceptance rules differ by jurisdiction.
How Certification Strengthens Product Quality And Patient Safety
Medical device suppliers may be several contractual steps removed from the patient, but their decisions can still affect device performance. A dimensional error, contaminated material, incomplete sterilization record, mislabeled package, or uncontrolled software change can create consequences far beyond the supplier’s facility.
Consistency Through Controlled Processes
ISO 13485 requires organizations to plan and control the processes that affect product quality. This includes establishing acceptance criteria, defining responsibilities, maintaining suitable work environments, controlling equipment, and retaining records that show what occurred.
Process control helps answer practical questions such as:
- Which approved instructions did employees follow?
- Were operators appropriately trained?
- Was the correct material used?
- Was measuring equipment calibrated?
- Were required inspections completed?
- Did the process remain within validated parameters?
- Who released the product?
- Can the affected lot or batch be identified?
Reliable answers are essential when a customer investigates a defect or determines whether additional products could be affected.
Validation Of Special Processes
Some process outcomes cannot be fully verified by inspecting the finished product. Sterilization, sterile-barrier sealing, welding, molding, coating, automated assembly, and certain software-controlled processes may require validation.
Validation uses documented evidence to show that a process can consistently produce results meeting predetermined requirements. The supplier must also maintain the validated state through equipment controls, monitoring, trained personnel, change evaluation, and revalidation when appropriate.
This approach is more protective than depending exclusively on end-of-line testing, particularly when testing every unit would be destructive, impractical, or unable to reveal latent defects.
Risk-Based Control Throughout Operations
ISO 13485 requires a risk-based approach to controlling appropriate quality management system processes. Risk influences how extensively an organization qualifies suppliers, validates processes, investigates problems, controls changes, and monitors performance.
A supplier should therefore avoid applying the same level of oversight to every item without considering its intended use or potential effect. A cosmetic packaging component and a critical implantable-device component do not necessarily warrant identical controls.
Risk-based decisions should be documented and supported by objective evidence. They should also be revisited when new information becomes available, including complaints, test failures, process changes, regulatory actions, or customer feedback.
Connecting Quality Records To Safety Decisions
Strong documentation enables faster, more accurate decisions when a problem emerges. If a manufacturer receives a complaint, supplier records may help determine:
- Whether the problem is isolated or systemic
- Which lots, batches, or serial numbers could be affected
- Whether a material or process change occurred
- Whether other customers received similar products
- Whether containment, correction, or field action may be necessary
The quality system therefore supports patient safety not merely by trying to prevent defects, but also by enabling effective responses when problems occur.
Why ISO 13485 Matters During Supplier Qualification
Medical device manufacturers are expected to evaluate and control suppliers according to the products and services they provide. ISO 13485 certification gives prospective customers a recognized starting point for that evaluation.
Demonstrating Organizational Maturity
A current, appropriately scoped certificate indicates that an independent body has assessed the supplier’s quality management system. This can provide initial confidence that the organization has established fundamental controls and submits to continuing external oversight.
Certification may help demonstrate that the supplier has defined processes for:
- Reviewing customer requirements
- Approving and monitoring its own vendors
- Controlling drawings and specifications
- Preserving products during handling and storage
- Managing nonconforming output
- Evaluating process or material changes
- Investigating complaints
- Performing corrective action
- Maintaining records
For customers comparing otherwise similar vendors, these controls can become an important differentiator.
Reducing—But Not Eliminating—Customer Audits
A credible ISO 13485 certificate may reduce the amount of basic quality-system verification a customer needs to perform. Buyers can focus more closely on product-specific risks, technical capabilities, capacity, special processes, and contractual responsibilities.
However, certification should not be the only qualification criterion. A medical device manufacturer may still need to conduct an on-site or remote audit, particularly when the supplier provides a critical component or high-risk service.
Customer evaluation may also include:
- Technical expertise and equipment
- Regulatory history
- Complaint or recall involvement
- Capacity and business continuity
- Cybersecurity and data controls
- Process capability
- Subsupplier management
- Delivery performance
- Financial stability
- Responsiveness to corrective actions
ISO 13485 certification supports due diligence; it does not replace it.
Establishing Clear Quality Agreements
Quality agreements define how the customer and supplier will manage activities affecting compliance and product quality. They are particularly valuable when responsibilities could otherwise be misunderstood.
An effective agreement may address:
- Specifications and acceptance criteria
- Required records and retention periods
- Audit rights
- Complaint investigation support
- Change-notification requirements
- Nonconformance handling
- Corrective-action timelines
- Traceability expectations
- Regulatory inspection notification
- Subcontracting restrictions
- Recall or field-action support
An ISO 13485 quality system gives these commitments an operational structure. Without functioning procedures and accountable personnel, even a well-written agreement may fail in practice.
How ISO 13485 Improves Traceability And Supply-Chain Control
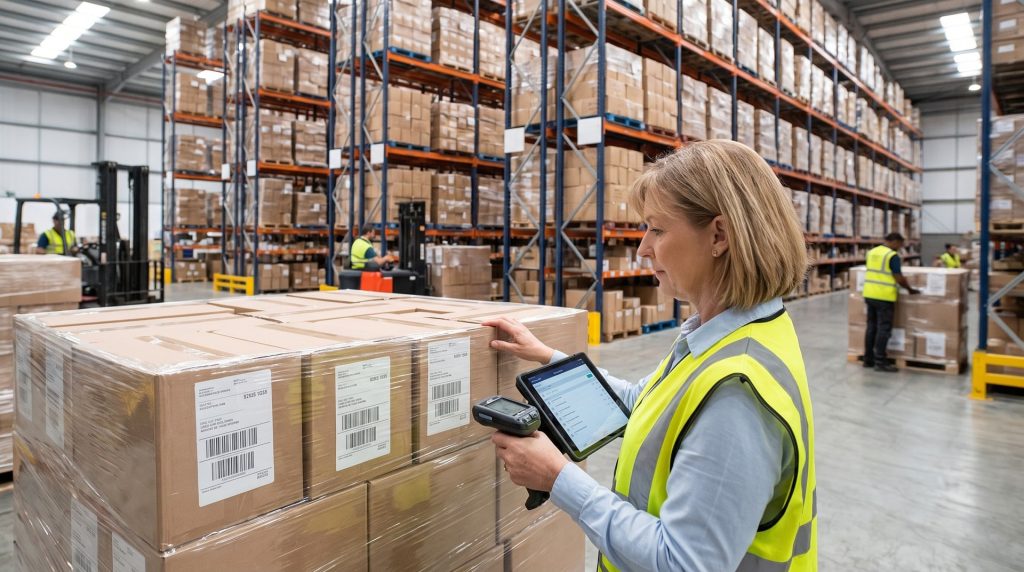
Medical device supply chains often include multiple material providers, contract manufacturers, testing laboratories, sterilizers, packagers, warehouses, and distributors. Each handoff can introduce risk unless responsibilities and records are controlled.
Traceability From Materials To Delivery
Traceability allows an organization to connect incoming materials, production activities, inspection results, release decisions, and distribution information. The required level of detail varies according to the device, its classification, regulatory requirements, and customer specifications.
A robust system may connect:
- Supplier name and material lot
- Purchase order and receipt date
- Inspection or test results
- Production lot or work order
- Equipment and process parameters
- Operators and inspectors
- Packaging and labeling records
- Release authorization
- Shipment destination
This information can help a manufacturer limit the scope of an investigation or recall. Weak records may force an organization to treat a much larger quantity of product as potentially affected.
Control Of Subsuppliers
A certified supplier may rely on other organizations for raw materials, calibration, processing, sterilization, logistics, or testing. Outsourcing the work does not eliminate the need to control it.
ISO 13485 requires appropriate evaluation, selection, monitoring, and reevaluation of suppliers. The extent of control should reflect the potential effect of the purchased product or service on subsequent product realization or the final medical device.
Important controls can include:
- Approved-supplier criteria
- Initial qualification
- Quality and technical agreements
- Performance metrics
- Certificate and accreditation verification
- Incoming verification
- Periodic reevaluation
- Change-notification obligations
- Escalation and disqualification procedures
These controls help prevent hidden weaknesses deeper in the supply chain.
Change Control And Notification
An apparently minor change can alter device performance. Examples include replacing a resin, moving production to another facility, modifying a test method, changing software, using a new sterilization load configuration, or switching a critical subsupplier.
A mature quality system requires changes to be evaluated, documented, approved, implemented, and verified in a controlled manner. Customer or regulatory approval may be required before implementation.
ISO 13485 certification provides evidence that a change-control process exists, but customers should still specify which changes require advance notification or written authorization.
Business Benefits For Medical Device Suppliers
ISO 13485 is fundamentally a quality and regulatory standard, but its disciplined controls can also produce meaningful commercial and operational benefits.
Access To More Customer Opportunities
Many medical device manufacturers include ISO 13485 certification in supplier questionnaires, requests for proposals, and purchasing requirements. Some will not consider an uncertified supplier for critical work unless a justified exception and enhanced oversight are available.
Certification can therefore help a supplier:
- Qualify for medical device contracts
- Respond more effectively to customer audits
- Demonstrate readiness for regulated work
- Compete across multiple geographic markets
- Build confidence with new business partners
- Support customers’ regulatory submissions and inspections
The actual benefit depends on the certificate’s scope and the supplier’s ability to demonstrate effective implementation.
Fewer Errors And More Predictable Operations
Documented procedures and defined controls can reduce dependence on informal knowledge. When employees know which instructions apply and records are consistently maintained, operations become easier to repeat and evaluate.
A strong quality system can help reduce:
- Use of obsolete drawings or procedures
- Unapproved material substitutions
- Missed inspections
- Incomplete batch records
- Undocumented process changes
- Recurring nonconformities
- Delayed complaint investigations
- Confusion over responsibilities
Reducing these failures can lower scrap, rework, returns, customer disruption, and investigation costs. Certification does not guarantee these outcomes, but it provides a framework for achieving them.
Better Corrective And Preventive Action
Corrective and preventive action, commonly called CAPA, is central to an effective medical device quality system. When a problem occurs, the organization should not simply repair the immediate defect. It should determine whether the issue is systemic, identify appropriate causes, implement proportionate action, and verify effectiveness.
Useful sources of quality data include:
- Internal audit findings
- Customer complaints
- Returned products
- Supplier performance
- Process deviations
- Nonconforming materials
- Trend reports
- Service records
- Regulatory observations
When CAPA is effective, recurring problems should decrease. When CAPA becomes a paperwork exercise, the same issue may return under a different lot number or product line.
Stronger Management Visibility
ISO 13485 requires management involvement and periodic review of quality-system performance. Leadership should receive meaningful information about complaints, audit results, process performance, supplier issues, CAPA, regulatory changes, resource needs, and improvement opportunities.
This prevents quality from becoming the responsibility of one department alone. Senior management remains accountable for ensuring that the system is resourced, maintained, and capable of meeting applicable requirements.
What Buyers Should Verify Beyond The Certificate
A certificate is valuable evidence, but it is only one part of responsible supplier assessment. Medical device buyers should verify whether the system is relevant, current, and effective.
Review The Scope And Authenticity
The certificate’s scope should match the supplier’s actual role. For example, certification for warehousing and distribution does not necessarily cover device design, sterile packaging, or contract manufacturing.
A customer should verify:
Verification Point | Why It Matters |
Certified legal entity | Confirms the certificate belongs to the contracting organization |
Covered address or site | Ensures the facility performing the work is included |
Certification scope | Shows which products, services, or activities were audited |
Issue and expiration dates | Helps confirm current validity |
Certification body | Identifies who conducted the assessment |
Accreditation information | Helps evaluate certification-body recognition |
Current certificate status | Detects suspension, withdrawal, or outdated documentation |
When appropriate, buyers can confirm certificate status directly through the issuing certification body or a recognized certification database.
Evaluate Practical Implementation
A supplier can have extensive procedures yet implement them poorly. Buyers should look for objective evidence that the system functions in daily operations.
Relevant evidence may include:
- Completed training records
- Current work instructions
- Lot and batch history records
- Validation reports
- Calibration records
- Supplier evaluations
- Nonconformance investigations
- CAPA effectiveness checks
- Internal audit results
- Management review outputs
The goal is not to collect documents unnecessarily. It is to verify that controls proportionate to the product’s risk are operating as intended.
Define Responsibilities Before Work Begins
The customer and supplier should agree on who is responsible for design decisions, risk-management inputs, purchasing controls, validation, final release, complaint investigation, regulatory reporting support, and change approval.
These responsibilities should appear in contracts, specifications, purchase controls, and quality agreements where appropriate. Ambiguous responsibility can delay investigations and create compliance gaps when urgent action is needed.
Monitor The Relationship Over Time
Supplier qualification is not a one-time event. Performance should be monitored using criteria that reflect the supplier’s risk and role.
Possible measures include:
- On-time delivery
- Incoming acceptance rates
- Nonconformance frequency
- Complaint involvement
- CAPA response time
- Audit findings
- Change-notification performance
- Documentation accuracy
- Recurring defects
- Certificate status
Performance trends may justify increased oversight, corrective action, requalification, or a change in approved-supplier status.
Conclusion
ISO 13485 certification is important for medical device suppliers because it provides internationally recognized evidence of a structured, independently assessed quality management system. It helps organizations control processes, document decisions, manage risk, maintain traceability, investigate problems, and support the regulatory responsibilities of their customers.
The standard is increasingly relevant to suppliers serving the United States because the FDA’s QMSR now incorporates ISO 13485:2016 by reference. It also provides a useful foundation for organizations operating across other regulated markets. Nevertheless, certification is not FDA approval, product certification, or a substitute for compliance with applicable national laws.
For buyers, an ISO 13485 certificate can simplify initial supplier evaluation, but it should be examined for validity, scope, site coverage, and relevance. Product-specific capabilities, quality records, regulatory history, change controls, and ongoing performance must also be considered.
For suppliers, the greatest value comes from using the quality management system as an active operating framework. When procedures accurately reflect the work, employees understand their responsibilities, management responds to quality data, and problems lead to effective corrective action, ISO 13485 can support safer products, stronger customer relationships, and more dependable supply chains.
Merit Pharmaceutical recognizes that medical device sourcing requires more than confirming product availability. Supplier qualifications, documentation, traceability, storage conditions, and regulatory considerations can all influence purchasing decisions. Working with knowledgeable suppliers helps healthcare and business customers select medical devices with greater confidence and appropriate attention to quality requirements.
Frequently Asked Questions
1. Is ISO 13485 Certification Legally Required For Every Medical Device Supplier?
No. Whether certification is legally required depends on the organization’s role, products, services, target markets, and applicable regulations. Some customers or commercial contracts may require certification even when a regulation does not explicitly require the supplier to hold a certificate. Organizations should evaluate the rules applying to each market and activity.
2. Does ISO 13485 Certification Mean A Medical Device Is FDA-Approved?
No. ISO 13485 certification assesses an organization’s quality management system. It does not provide FDA clearance, approval, registration, listing, or authorization to market a device. Those are separate regulatory matters determined by the device and its applicable requirements.
3. What Is The Relationship Between ISO 13485 And The FDA’s QMSR?
The FDA’s QMSR, effective February 2, 2026, incorporates ISO 13485:2016 by reference as the foundation of U.S. medical device quality-system requirements. Manufacturers must also comply with applicable provisions of the Federal Food, Drug, and Cosmetic Act and other FDA regulations. An ISO certificate does not prevent FDA inspection or independently prove full U.S. compliance.
4. Does ISO 13485 Certification Apply Only To Finished-Device Manufacturers?
No. The standard may be used by organizations throughout the medical device life cycle, including component manufacturers, contract manufacturers, sterilization providers, testing laboratories, packaging companies, distributors, and service providers. The certification scope should accurately identify the activities covered.
5. How Can A Buyer Verify An ISO 13485 Certificate?
The buyer should review the company name, facility address, certification scope, issuing certification body, certificate number, and validity dates. When the supplier’s role is critical, the buyer may also verify the certificate through the issuing body and review supporting quality records or conduct an audit.
6. Can ISO 13485 Certification Replace A Supplier Audit?
Not always. Certification may reduce the need to audit basic quality-system elements, but a customer may still need to examine product-specific capabilities, special processes, validation, capacity, change controls, or regulatory risks. The depth of supplier oversight should be proportionate to the supplied product or service.
7. How Often Must An ISO 13485-Certified Supplier Be Audited?
Certification programs generally include periodic surveillance audits and recertification assessments according to the certification body’s audit program. Customers may conduct their own audits based on supplier risk, performance, contract terms, quality issues, or significant operational changes. Certification and customer auditing serve related but distinct purposes.